
| Jung-Hwan Kim | 4 Articles |
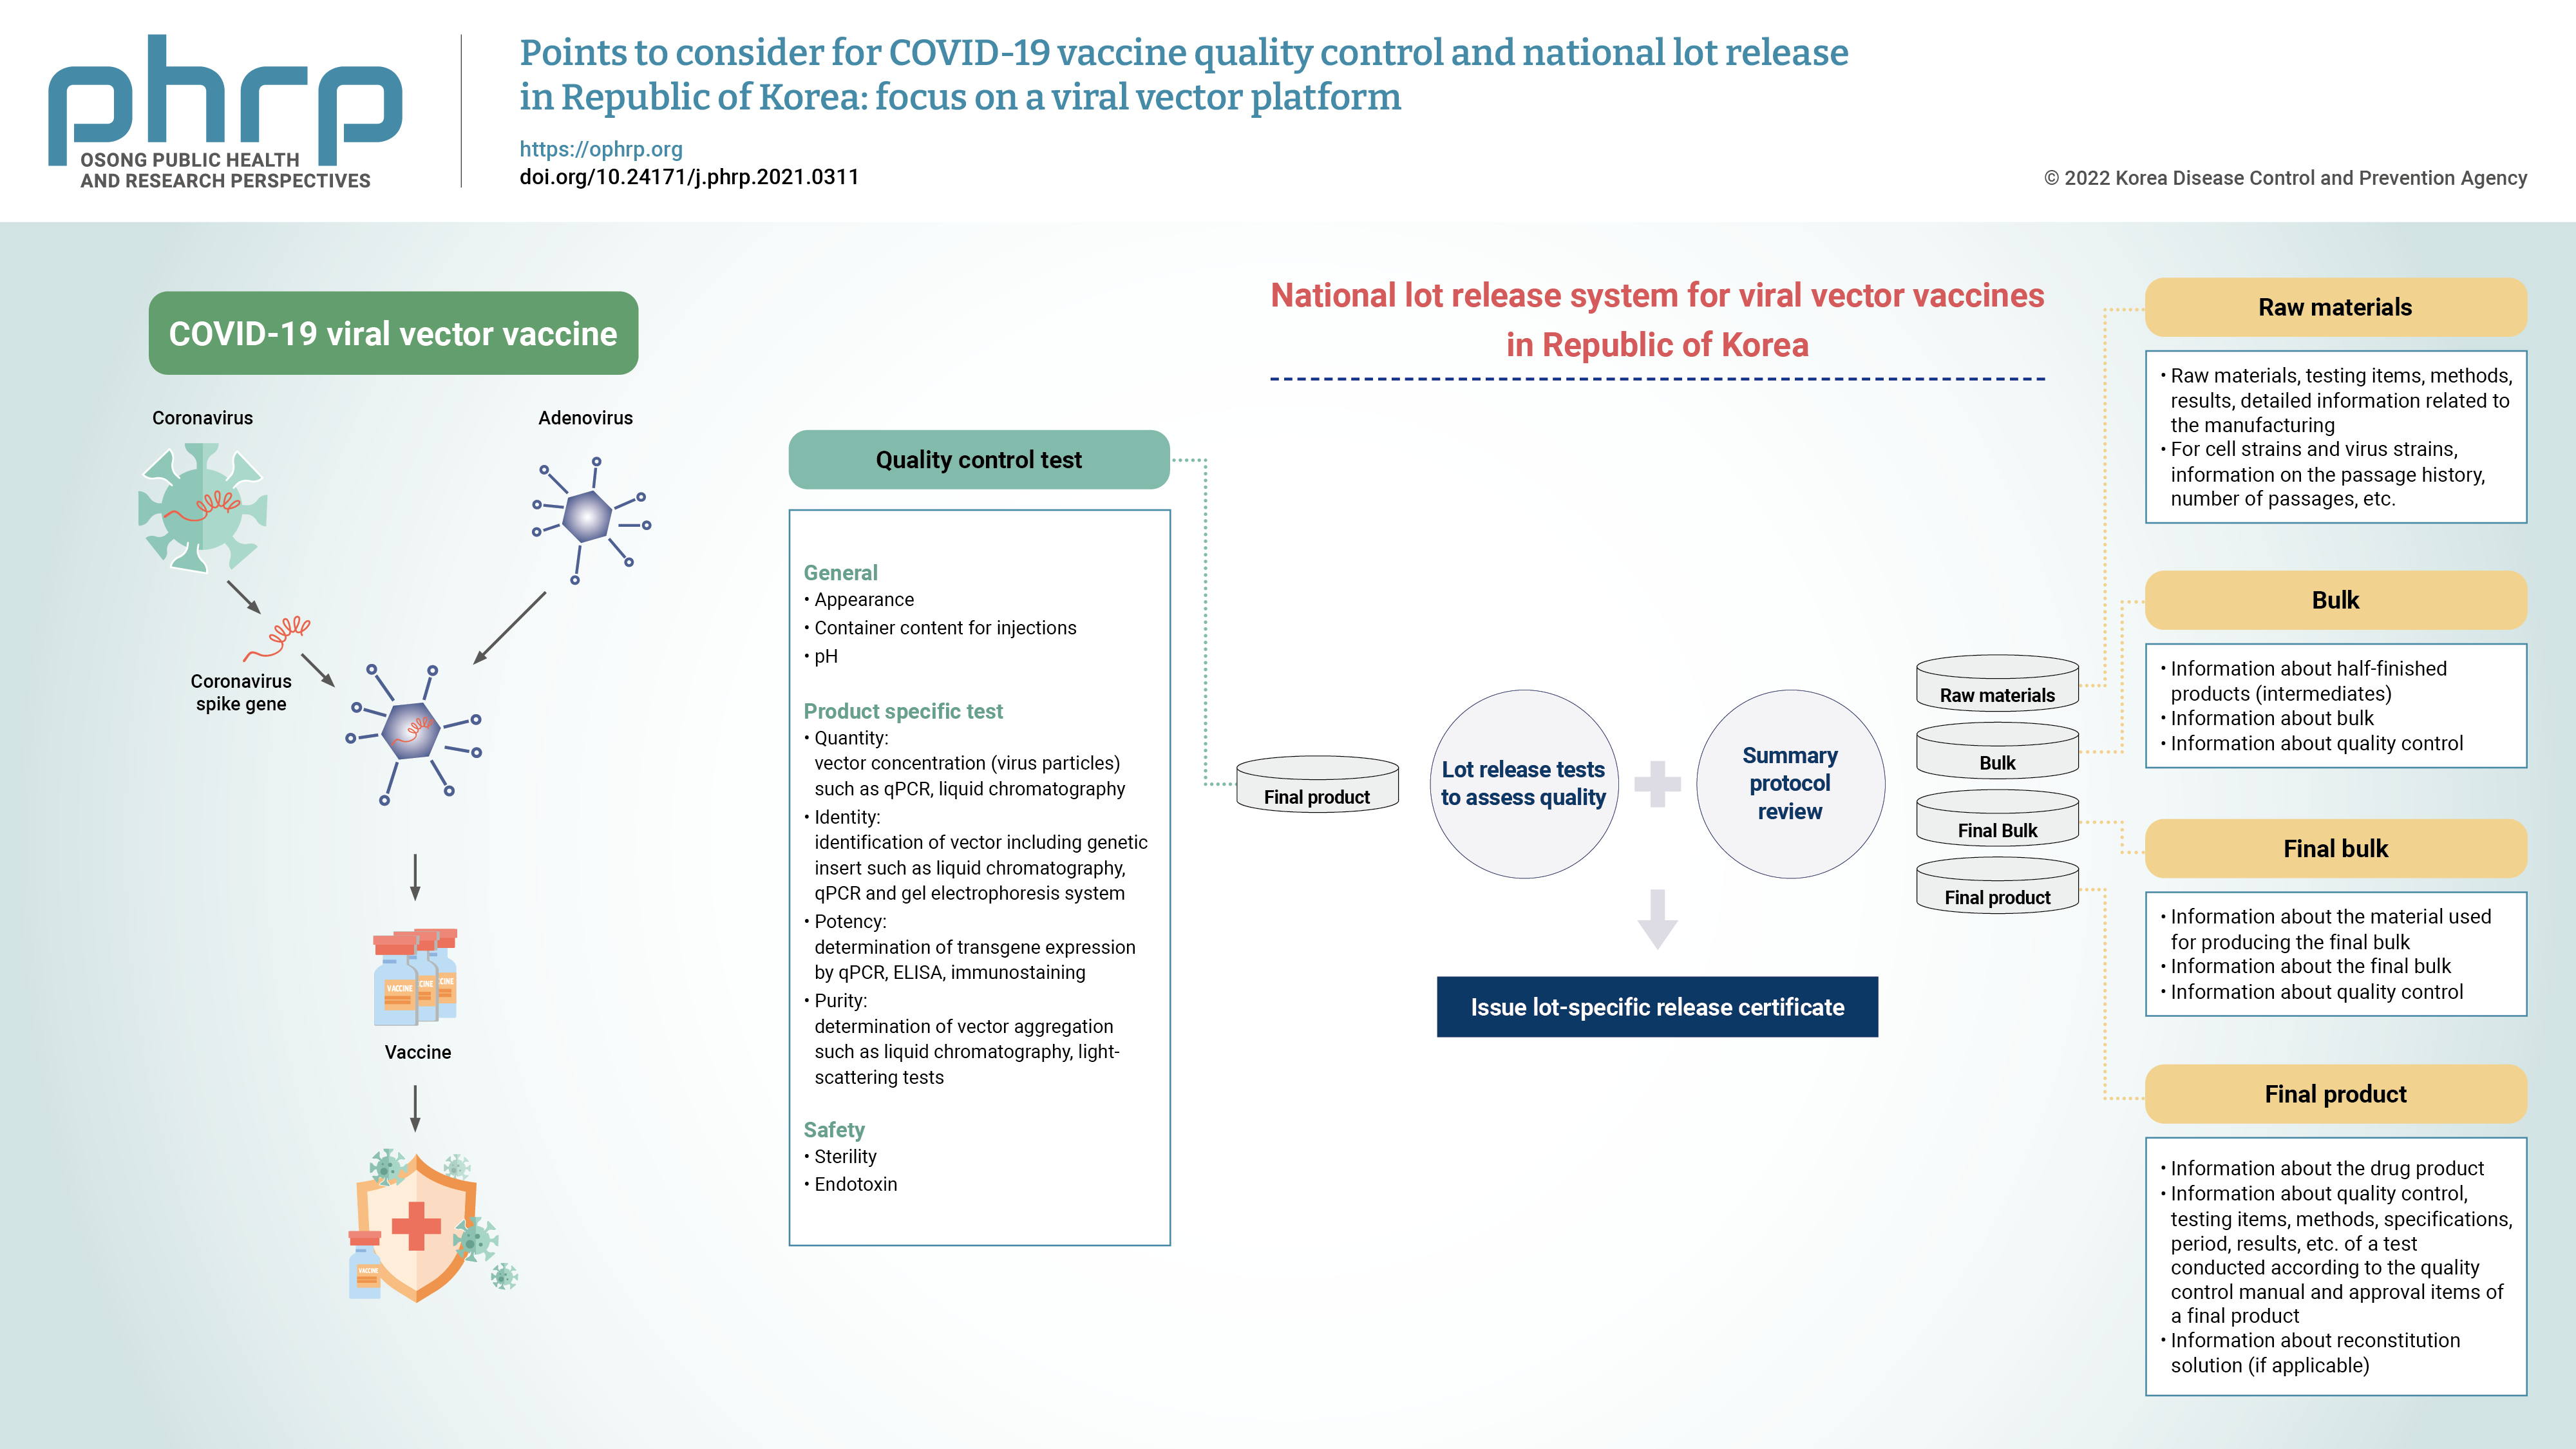
Due to the global public health crisis caused by the coronavirus disease 2019 (COVID-19) pandemic, the importance of vaccine development has increased. In particular, a rapid supply of vaccines and prompt deployment of vaccination programs are essential to prevent and overcome the spread of COVID-19. As a part of the vaccine regulations, national lot release is regulated by the responsible authorities, and this process involves the assessment of the lot before a vaccine is marketed. A lot can be released for use when both summary protocol (SP) review and quality control testing are complete. Accelerated lot release is required to distribute COVID-19 vaccines in a timely manner. In order to expedite the process by simultaneously undertaking the verification of quality assessment and application for approval, it is necessary to prepare the test methods before marketing authorization. With the prolonged pandemic and controversies regarding the effectiveness of the COVID-19 vaccine against new variants, public interest for the development of a new vaccine are increasing. Domestic developers have raised the need to establish standard guidance on the requirements for developing COVID-19 vaccine. This paper presents considerations for quality control in the manufacturing process, test items, and SP content of viral vector vaccines.
Citations
Citations to this article as recorded by

<sec>
<title>Objectives</title>
<p>Vaccinations against diphtheria and tetanus are essential in providing immunity against these bacterial infections. The potency of diphtheria and tetanus toxoid vaccines can be measured using the in vivo toxin neutralization assay. The limit of potency of this assay was determined only for children. Therefore, we assessed the potency of adult vaccines using this assay to identify the feasibility of limit for adult vaccines.</p></sec>
<sec>
<title>Methods</title>
<p>Fifteen lots of tetanus-reduced diphtheria and tetanus-diphtheria-acellular pertussis vaccines were used. In vivo toxin neutralization and lethal challenge assays were conducted on each vaccine to calculate the potencies of the toxoids. National reference standards for toxins and antitoxins were used for in vivo toxin neutralization assay.</p></sec>
<sec>
<title>Results</title>
<p>All 15 lots satisfied the limits of potency for lethal challenge assay. The potency of diphtheria and tetanus toxoids exceeded 1 and 8 units/mL, respectively, for in vivo toxin neutralization assay.</p></sec>
<sec>
<title>Conclusion</title>
<p>Although additional studies are required for new assays and limits, the current level of potency for adult vaccines as determined by in vivo toxin neutralization assay, was demonstrated in this study. Such efforts to improve assays are expected to promote the development of diphtheria and tetanus vaccines for adults and to contribute to vaccine self-sufficiency.</p></sec>
Citations
Citations to this article as recorded by
Objectives
To develop and optimize quantitative HPLC method using 2,3-naphthalenedicarboxaldehyde (NDA) after simple and efficient solid phase extraction to determine the histamine in a biopharmaceutical (Histobulin™). Methods The HPLC method was established using NDA-induced Histobulin and compared with the recently reported HPLC method using o-phthaldehyde (OPA). The validated NDA-applied HPLC method was adjusted to 15 lots of Histobulin and compared by the current lot-release-test method using fluorimetry in recovery of histamine and reproducibility. Results Analyses of six HPLC chromatograms using NDA and OPA each were compared. NDA produced a more stable chromatogram baseline than OPA, and showed better stability. The HPLC analysis was validated in accuracy (91–103%), precision (interday/intraday assay CV ≤2.30%), and linearity of dose–response curve (R<sup>2</sup> ≥ 0.9919). The detection limit was 0.0076 μg/mL and the quantitative limit was 0.0229 μg/mL. The amount of histamine per 12 mg of immunoglobulin was determined to be 0.17 ± 0.016 μg by the HPLC and 0.025 ± 0.013 μg by the current lot-release-test method using fluorimetry. Conclusion NDA derivatization showed better stability compared with the OPA method. Therefore the newly established NDA-derivatizated HPLC method may be more suitable than the fluorimetric method in lot-release-tests of biopharmaceuticals. Citations
Citations to this article as recorded by
|
|


